
In pharmaceutical development, the ultimate goal is not simply to manufacture a tablet or capsule that meets specification, but to consistently deliver the intended therapeutic effect to patients. Among the analytical tools available to scientists, dissolution testing remains one of the most powerful techniques for understanding how a drug product is likely to behave once administered.
Historically, dissolution testing was viewed primarily as a quality control requirement, used to demonstrate batch-to-batch consistency during manufacturing. Today, however, the role of dissolution has expanded significantly. Regulatory agencies increasingly recognise dissolution testing as a valuable tool for understanding product performance, supporting formulation development, establishing clinically relevant specifications, and even predicting in vivo behaviour.
As pharmaceutical products become more complex and development timelines become increasingly compressed, incorporating dissolution testing early in development has become an essential strategy for reducing risk and accelerating product progression.
What is Dissolution Testing?
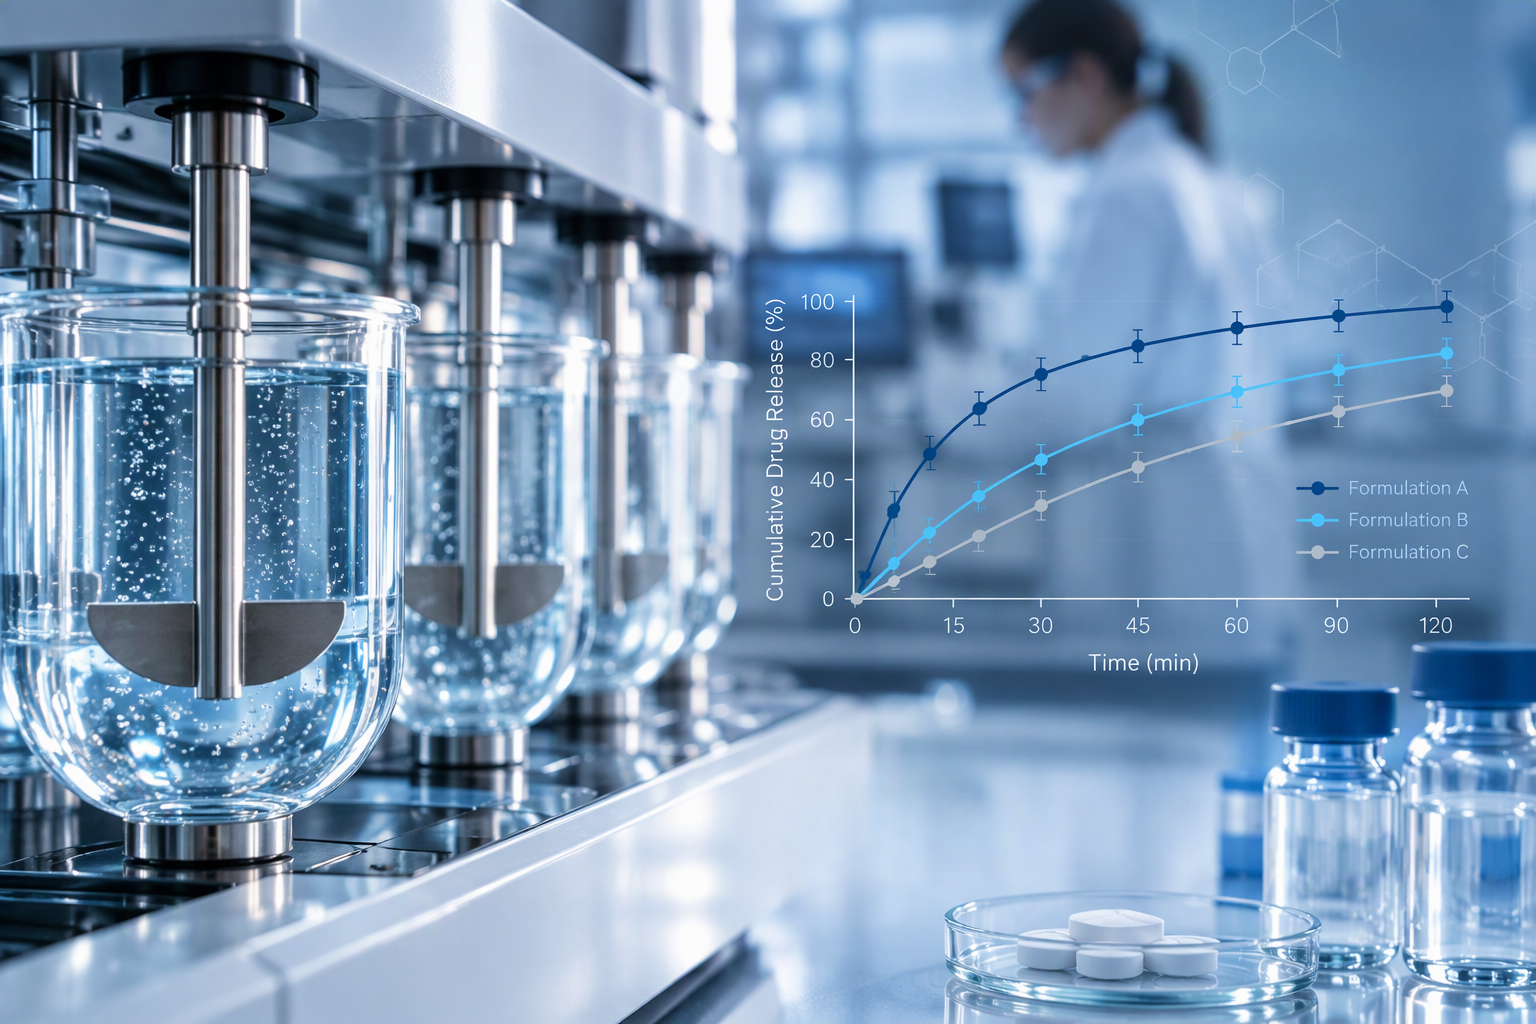
Dissolution testing measures the rate and extent to which an active pharmaceutical ingredient (API) is released from a dosage form into a dissolution medium under controlled laboratory conditions.
For oral solid dosage forms such as tablets and capsules, dissolution is often one of the most critical steps that determines drug absorption. If a drug is not released appropriately from its formulation, it cannot become available for absorption across the gastrointestinal tract, regardless of the potency of the active ingredient.
Using standardised apparatus described within the United States Pharmacopeia (USP) and European Pharmacopoeia (Ph. Eur.), dissolution profiles can be generated that characterise product performance across a range
of physiological conditions. These data provide valuable insight into formulation robustness, manufacturing consistency, and potential bioavailability.
Dissolution Beyond Quality Control
While dissolution testing remains a cornerstone of quality control, its value extends far beyond routine release testing.
During formulation development, dissolution testing enables scientists to:
- Compare prototype formulations.
- Assess the impact of excipient selection.
- Investigate particle size effects.
- Evaluate manufacturing process changes.
- Support formulation optimisation.
- Identify potential performance risks before clinical studies.
Early dissolution investigations can reveal significant differences between formulations that may not be evident from conventional physicochemical testing alone. This information can guide development decisions before substantial resources have been committed.
For example, changes in granulation methodology, compression force, or excipient grade may have little impact on assay or content uniformity results while significantly affecting dissolution performance. Identifying such effects early can prevent costly delays during later development stages.
Growing Regulatory Focus on Dissolution
Regulatory expectations around dissolution testing have evolved considerably over the past decade.
Authorities including the U.S. Food and Drug Administration (FDA), European Medicines Agency (EMA), and International Council for Harmonisation (ICH) increasingly promote science-based approaches that utilise dissolution data throughout the product lifecycle.
A key area of focus has been the development of clinically relevant dissolution methods and specifications. Rather than demonstrating only that a product passes an arbitrary dissolution criterion, regulators now encourage the use of dissolution methods that discriminate between meaningful formulation or process changes and that provide insight into potential in vivo performance.
The emergence of Quality by Design (QbD) frameworks has further strengthened the role of dissolution testing. Within QbD, dissolution is frequently identified as a critical quality attribute (CQA) and is often used to assess the impact of critical material attributes (CMAs) and critical process parameters (CPPs).
Consequently, dissolution data are increasingly used to support:
- Formulation selection.
- Manufacturing process development.
- Control strategy establishment.
- Specification setting.
- Regulatory submissions.
- Post-approval change management.
Organisations that develop scientifically justified dissolution methods early in development are often better positioned to navigate regulatory interactions and product lifecycle challenges.
Dissolution as a Biopredictive Tool
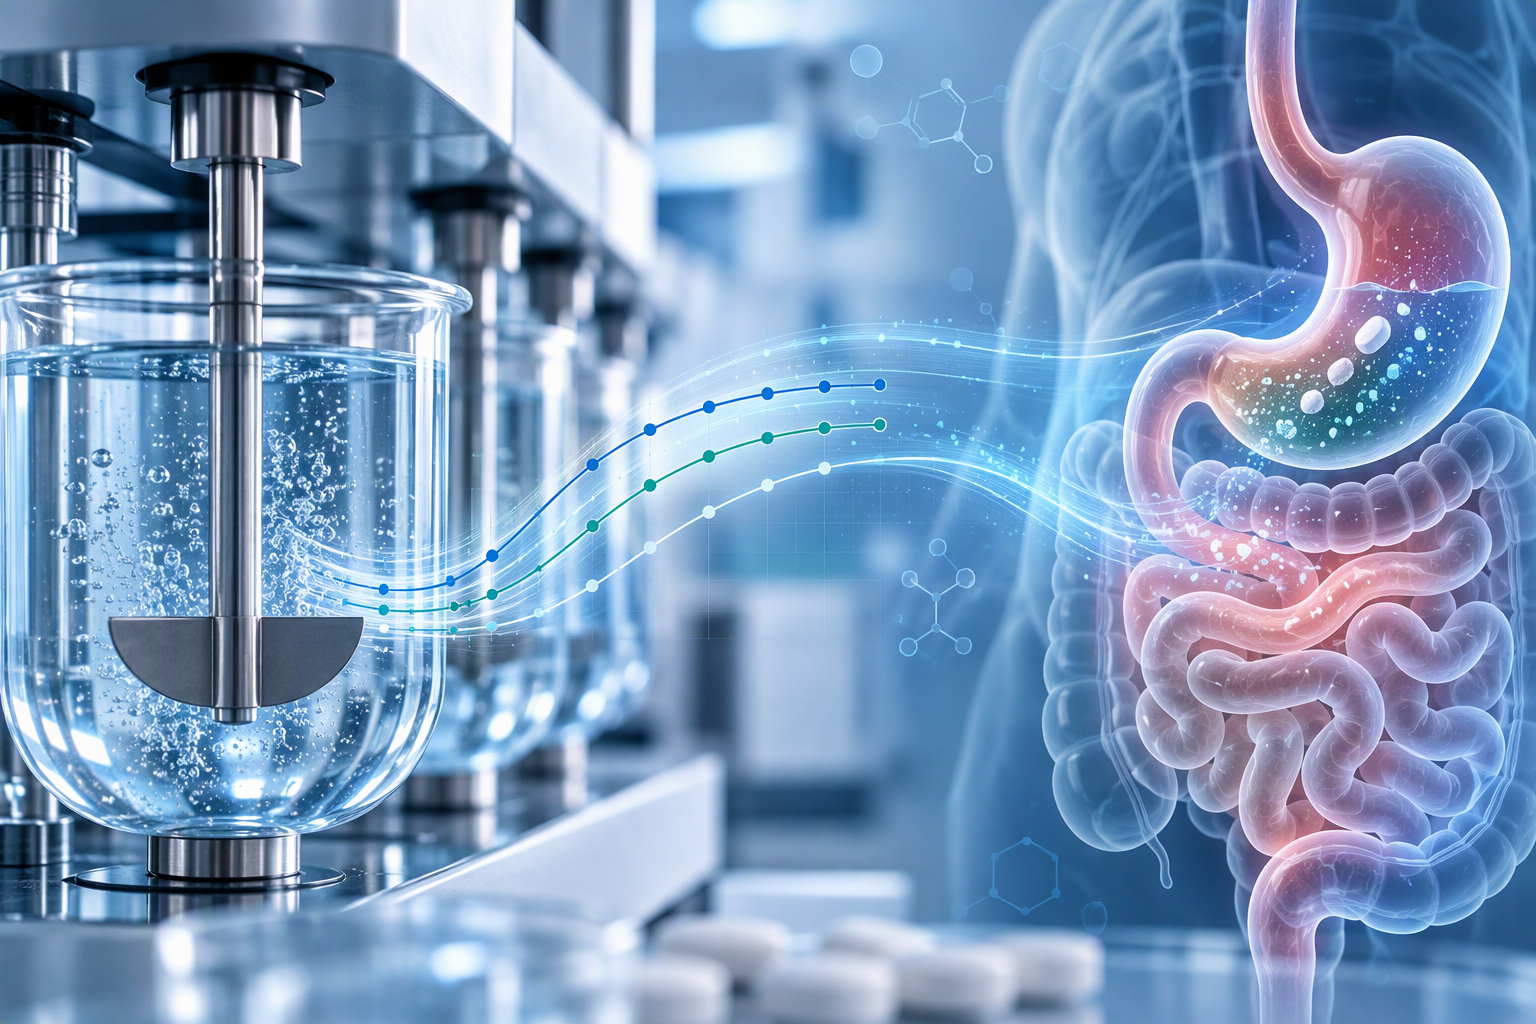
One of the most significant developments in pharmaceutical science has been the growing emphasis on biopredictive dissolution testing.
Traditional quality control dissolution methods are designed primarily for reproducibility and robustness. Although these methods are essential for routine testing, they do not always reflect conditions encountered within the human gastrointestinal tract.
Biopredictive dissolution methods seek to bridge this gap by incorporating physiologically relevant conditions, including:
- Biorelevant media compositions.
- Dynamic pH environments.
- Physiologically representative agitation conditions.
- Fed and fasted state assessments.
- Absorption-related considerations.
When combined with physiologically based pharmacokinetic (PBPK) modelling, dissolution data can contribute to a more comprehensive understanding of in vivo drug performance.
This approach is particularly valuable for:
- Poorly soluble compounds.
- Modified release formulations.
- Complex generic products.
- Drug products seeking biowaiver opportunities.
- Formulations with known absorption limitations.
As regulators continue to promote model-informed drug development approaches, high-quality dissolution data are becoming increasingly important in demonstrating product understanding and supporting regulatory decision-making.
Why Early Dissolution Assessment Creates Value
Despite the recognised importance of dissolution testing, many development programmes delay comprehensive dissolution assessment until later stages.
This approach can create significant challenges.
Method development conducted late in the programme often encounters issues such as:
- Limited discriminatory power.
- Excessive method variability.
- Inadequate understanding of formulation behaviour.
- Difficulty establishing clinically relevant specifications.
- Increased regulatory scrutiny.
By contrast, early dissolution assessment enables development teams to build a stronger scientific foundation from the outset.
Conducting exploratory dissolution studies during formulation screening can help identify performance trends, understand release mechanisms, and establish initial method parameters. As development progresses, this knowledge can be leveraged to create robust methods that remain fit for purpose throughout the product lifecycle.
Early assessment also allows organisations to generate valuable historical data that can support future investigations, specification setting activities, and regulatory interactions.
Perhaps most importantly, early dissolution studies can identify formulation liabilities before significant investment has been made in scale-up, validation, and regulatory submissions.
Supporting Smarter Development Decisions
In today’s pharmaceutical environment, dissolution testing should not be viewed merely as a regulatory obligation. Instead, it should be regarded as a strategic development tool capable of generating critical insights into product performance.
The industry’s increasing focus on clinically relevant dissolution, Quality by Design principles, and model-informed drug development ensures that dissolution testing will continue to play an expanding role throughout the pharmaceutical lifecycle.
For development organisations seeking to accelerate programmes, reduce technical risk, and strengthen regulatory confidence, incorporating dissolution assessment at the earliest stages of formulation development can provide a significant advantage.
At Biowaived, we believe that dissolution testing is most valuable when applied proactively. Early-stage dissolution assessment can help developers understand formulation behaviour sooner, support informed development decisions, and establish `the foundations for robust, scientifically justified dissolution methods that remain effective throughout product commercialisation.